|
|
| (3 intermediate revisions by 3 users not shown) |
| Line 1: |
Line 1: |
| {{Infobox script-repo | | {{Infobox script-repo |
| |type = module | | |type = module |
| |filename = findSurfaceResidues.py | | |filename = scripts/findSurfaceResidues.py |
| |author = [[User:inchoate|Jason Vertrees]] | | |author = [[User:inchoate|Jason Vertrees]] |
| |license = BSD-2-Clause | | |license = BSD-2-Clause |
| }} | | }} |
|
| |
|
| <gallery>
| | The [[findSurfaceResidues]] script will select (and color if requested) surface residues and atoms on an object or selection. See the options below. |
| Image:FindExRes.png|thumb|right|300px|Result of $TUT/1hpv.pdb at 2.5 Ang cutoff.
| |
| Image:Surface_residues_ex.png|300px|Example coloring of surface residues
| |
| </gallery>
| |
|
| |
|
| | Each time, the script will create two new selections called, '''exposed_res_XYZ''' and '''exposed_atm_XYZ''' where ''XYZ'' is some random number. This is done so that no other selections/objects are overwritten. |
|
| |
|
| = Overview = | | = Usage = |
| This script will select (and color if requested) surface residues and atoms on an object or selection. See the options below.
| |
|
| |
|
| | findSurfaceResidues [ selection=all [, cutoff=2.5 [, doShow=0 ]]] |
|
| |
|
| Each time, the script will create two new selections called, 'exposed_res_XYZ' and 'exposed_atm_XYZ' where XYZ is some random number. This is done so that no other selections/objects are overwritten.
| | = Arguments = |
|
| |
|
| = Usage = | | * '''selection''' = str: The object or selection for which to find exposed residues {default: all} |
| | * '''cutoff''' = float: The cutoff in square Angstroms that defines exposed or not. Those atoms with > cutoff Ang^2 exposed will be considered ''exposed'' {default: 2.5 Ang^2} |
| | * '''doShow''' = 0/1: Change the visualization to highlight the exposed residues vs interior {default: 0} |
|
| |
|
| findSurfaceResidues [ selection=all [, cutoff=2.5 [, doShow=False ]]]
| | = Examples = |
|
| |
|
| The parameters are:
| | <gallery> |
| | Image:FindExRes.png|thumb|right|300px|Result of $TUT/1hpv.pdb at 2.5 Ang cutoff. |
| | Image:Surface_residues_ex.png|300px|Example coloring of surface residues |
| | </gallery> |
|
| |
|
| '''selection'''
| | <source lang="python"> |
| :: The object or selection for which to find exposed residues;
| | run findSurfaceResidues.py |
| :: DEFAULT = (all)
| | fetch 1hpv, async=0 |
| '''cutoff'''
| |
| :: The cutoff in square Angstroms that defines exposed or not. Those residues with > cutoff Ang^2 exposed will be considered ''exposed'';
| |
| :: DEFAULT = 2.5 Ang^2
| |
| '''doShow'''
| |
| :: Change the visualization to highlight the exposed residues vs interior
| |
| :: DEFAULT = False/Blank
| |
|
| |
|
| == Examples ==
| |
| <source lang="python">
| |
| # make sure you download and run the code below, before trying these examples.
| |
| load $TUT/1hpv.pdb
| |
| findSurfaceResidues | | findSurfaceResidues |
| | |
| # now show the exposed | | # now show the exposed |
| findSurfaceResidues doShow=True | | findSurfaceResidues doShow=1 |
|
| |
|
| # watch how the visualization changes: | | # watch how the visualization changes: |
| Line 51: |
Line 45: |
| </source> | | </source> |
|
| |
|
| = The Code = | | = See Also = |
| <source lang="python">
| |
| # -*- coding: utf-8 -*-
| |
| import pymol
| |
| from pymol import cmd
| |
| import random
| |
|
| |
|
| def findSurfaceResidues(objSel="(all)", cutoff=2.5, doShow=False, verbose=False):
| | * [[get_sasa_relative]] |
| """
| | * [[Get_Area|get_area]] |
| findSurfaceResidues
| |
| finds those residues on the surface of a protein
| |
| that have at least 'cutoff' exposed A**2 surface area.
| |
| | |
| PARAMS
| |
| objSel (string)
| |
| the object or selection in which to find
| |
| exposed residues
| |
| DEFAULT: (all)
| |
| | |
| cutoff (float)
| |
| your cutoff of what is exposed or not.
| |
| DEFAULT: 2.5 Ang**2
| |
| | |
| asSel (boolean)
| |
| make a selection out of the residues found
| |
| | |
| RETURNS
| |
| (list: (chain, resv ) )
| |
| A Python list of residue numbers corresponding
| |
| to those residues w/more exposure than the cutoff.
| |
| | |
| """
| |
| tmpObj="__tmp"
| |
| cmd.create( tmpObj, objSel + " and polymer");
| |
| if verbose!=False:
| |
| print "WARNING: I'm setting dot_solvent. You may not care for this."
| |
| cmd.set("dot_solvent");
| |
| cmd.get_area(selection=tmpObj, load_b=1)
| |
| | |
| # threshold on what one considers an "exposed" atom (in A**2):
| |
| cmd.remove( tmpObj + " and b < " + str(cutoff) )
| |
| | |
| stored.tmp_dict = {}
| |
| cmd.iterate(tmpObj, "stored.tmp_dict[(chain,resv)]=1")
| |
| exposed = stored.tmp_dict.keys()
| |
| exposed.sort()
| |
| | |
| randstr = str(random.randint(0,10000))
| |
| selName = "exposed_atm_" + randstr
| |
| if verbose!=False:
| |
| print "Exposed residues are selected in: " + selName
| |
| cmd.select(selName, objSel + " in " + tmpObj )
| |
| selNameRes = "exposed_res_" + randstr
| |
| cmd.select(selNameRes, "byres " + selName )
| |
| | |
| | |
| if doShow!=False:
| |
| cmd.show_as("spheres", objSel + " and poly")
| |
| cmd.color("white", objSel)
| |
| cmd.color("red", selName)
| |
| | |
| cmd.delete(tmpObj)
| |
| | |
| return exposed
| |
| | |
| | |
| cmd.extend("findSurfaceResidues", findSurfaceResidues)
| |
| </source>
| |
| | |
| = Another version =
| |
| <gallery heights="240px" widths="340px">
| |
| Image:Surfaceatoms.png|Another version, to show the same thing. The part of the protein which is not surface exposed is in cartoon. The white is the surface "byres" exposed residues shown in sticks. The orange is the atoms which is exposed.
| |
| </gallery>
| |
| | |
| <source lang="python">
| |
| from pymol import cmd, stored
| |
|
| |
| def surfaceatoms(molecule="NIL",show=True, verbose=True, cutoff=2.5):
| |
| """
| |
| surfaceatoms
| |
| finds those residues on the surface of a protein
| |
| that have at least 'cutoff' exposed A**2 surface area.
| |
| PARAMS
| |
| molecule (string)
| |
| the object or selection in which to find
| |
| exposed residues
| |
| DEFAULT: (last molecule in pymol)
| |
| cutoff (float)
| |
| your cutoff of what is exposed or not.
| |
| DEFAULT: 2.5 Ang**2
| |
| RETURNS
| |
| (list: (chain, resv ) )
| |
| A Python list of residue numbers corresponding
| |
| to those residues w/more exposure than the cutoff.
| |
| """
| |
| if molecule=="NIL":
| |
| assert len(cmd.get_names())!=0, "Did you forget to load a molecule? There are no objects in pymol."
| |
| molecule=cmd.get_names()[-1]
| |
| tmpObj="__tmp"
| |
| cmd.create(tmpObj, "(%s and polymer) and not resn HOH"%molecule)
| |
| if verbose!=False:
| |
| print "WARNING: I'm setting dot_solvent. You may not care for this."
| |
| cmd.set("dot_solvent")
| |
| cmd.get_area(selection=tmpObj, load_b=1)
| |
| # threshold on what one considers an "exposed" atom (in A**2):
| |
| cmd.remove( tmpObj + " and b < " + str(cutoff) )
| |
| stored.tmp_dict = {}
| |
| cmd.iterate(tmpObj, "stored.tmp_dict[(chain,resv)]=1")
| |
| exposed = stored.tmp_dict.keys()
| |
| exposed.sort()
| |
|
| |
| selName = "%s_atoms"%molecule
| |
| cmd.select(selName, molecule + " in " + tmpObj )
| |
| if verbose!=False:
| |
| print "Exposed residues are selected in: " + selName
| |
| selNameRes = "%s_resi"%molecule
| |
| cmd.select(selNameRes, "byres " + selName )
| |
|
| |
| if show!=False:
| |
| cmd.hide("everything", molecule)
| |
| cmd.show("cartoon", "%s and not %s and not resn HOH"%(molecule,selNameRes))
| |
| cmd.show("sticks", "%s"%selNameRes)
| |
| cmd.util.cbaw(selNameRes)
| |
| cmd.disable(selNameRes)
| |
| cmd.alter('%s'%(selName),'vdw=0.5')
| |
| cmd.show("spheres", "%s"%selName)
| |
| cmd.util.cbao(selName)
| |
| cmd.disable(selName)
| |
| | |
| cmd.delete(tmpObj)
| |
| print(exposed)
| |
| return(exposed)
| |
| cmd.extend("surfaceatoms", surfaceatoms)
| |
| </source>
| |
|
| |
|
| [[Category:Script_Library]] | | [[Category:Script_Library]] |
| Line 190: |
Line 54: |
| [[Category:Biochemical_Scripts]] | | [[Category:Biochemical_Scripts]] |
| [[Category:Structural_Biology_Scripts]] | | [[Category:Structural_Biology_Scripts]] |
| | [[Category:Pymol-script-repo]] |
| | __NOTOC__ |
The findSurfaceResidues script will select (and color if requested) surface residues and atoms on an object or selection. See the options below.
Each time, the script will create two new selections called, exposed_res_XYZ and exposed_atm_XYZ where XYZ is some random number. This is done so that no other selections/objects are overwritten.
Usage
findSurfaceResidues [ selection=all [, cutoff=2.5 [, doShow=0 ]]]
Arguments
- selection = str: The object or selection for which to find exposed residues {default: all}
- cutoff = float: The cutoff in square Angstroms that defines exposed or not. Those atoms with > cutoff Ang^2 exposed will be considered exposed {default: 2.5 Ang^2}
- doShow = 0/1: Change the visualization to highlight the exposed residues vs interior {default: 0}
Examples
run findSurfaceResidues.py
fetch 1hpv, async=0

findSurfaceResidues

# now show the exposed
findSurfaceResidues doShow=1

# watch how the visualization changes:
findSurfaceResidues doShow=1, cutoff=0.5
findSurfaceResidues doShow=1, cutoff=1.0
findSurfaceResidues doShow=1, cutoff=1.5
findSurfaceResidues doShow=1, cutoff=2.0
findSurfaceResidues doShow=1, cutoff=2.5
findSurfaceResidues doShow=1, cutoff=3.0
See Also
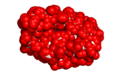 Result of $TUT/1hpv.pdb at 2.5 Ang cutoff.
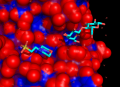 Example coloring of surface residues

